Chemistry Department Seminar

Electrochemistry: Application to Energy Storage and Critical Minerals

Abstract: Technologies like Li-ion battery (LIB) energy storage and beyond-Si semiconductor technologies at scales ranging from industrial installations to the phones in our pockets have become an increasingly important aspect of modern life. Their explosive growth, however, has led to new challenges to overcome. Increasing use of LIBs has led to concerns with material supply chains, battery safety and fundamental material-driven performance limitations. Similarly, the critical minerals (e.g., Ga, In) that are used in advanced semiconductor technologies also face supply chain concerns that must be addressed. Our group leverages fundamental and applied electrochemical research to help solve these and other challenges facing our nation. 

In my presentation, I will begin by discussing our research into a family of metal mixed chalcogenides, nickel sulfoselenides, and explore their use as electrocatalysts in rechargeable Zn-air batteries (ZABs). ZABs use earth-abundant and non-flammable materials to store energy, making them exciting for use in grid storage applications. Our work shows that controlling the ratio of S/Se in these materials influences their electrocatalytic performance, and thus allows for improved battery performance. In the second part of the talk, I will discuss our recent work in the development of an electrochemical method for recovering Ga from semiconductor manufacturing waste. I will discuss how electrochemical redox reactions can be used to replace strong acids in chemical leaching processes while simultaneously leveraging electrochemical specificity to selectively collect specific products from complex leachate solutions. Together, these projects provide a broad overview of the ways that electrochemistry can be used to support our modern world.

Biography: Dr. Bryan R. Wygant received his B.S. in chemistry from Texas Tech University in 2014 and his Ph.D. in chemistry from the University of Texas at Austin in 2020. At UT-Austin, he studied electrochemical materials for energy storage and production with Dr. C. Buddie Mullins, focusing on electrochemical water splitting and perovskite photovoltaics. After receiving his Ph.D., Wygant joined Sandia National Laboratories as a postdoctoral appointee, working with Dr. Timothy N. Lambert to study advanced cathode materials for batteries. Since becoming a senior member of technical staff at Sandia in 2023, Wygant has continued to explore how electrochemistry and materials can be used to solve important problems related to energy storage. His group explores such topics as earth-abundant electrocatalysts for alkaline Zn-air batteries, the safety of emerging Li-ion battery chemistries and advanced recycling techniques for battery and semiconductor materials, among other projects.
Date:
Location:
CP-114

Sustainable and Alternative Fuels: Turning Trash into Treasure with Heterogeneous Catalysis

Abstract: The growing need to substitute fossil energy sources with cleaner liquid transportation fuels has led to a proliferation of catalytic technologies that convert a huge variety of feedstocks into useful commodities. Many of these processes have been deployed on an industrial scale, but barriers to full adoption and widespread commercialization remain. 

These barriers include the expensive and unstable nature of catalyst materials, limited quality and or quantities of feedstock and the capital expenditures required to achieve economies of scale. One of the key ways in which these challenges may be overcome is through the development of improved catalysts- and the most effective means of enhancing catalyst performance is by better understanding the underlying chemistry. To that end, the sustainable and alternative fuels group at the UK Center for Applied Energy Research conducts research along three distinct but interrelated tracks that combine spectroscopic measurements, assessment of reaction performance and computational modelling to derive structure-activity relationships: 

  • The first track involves the development of catalytic materials to produce renewable diesel from fats oils and greases through the decarboxylation and decarbonylation reactions, a pathway that allows the use of inexpensive base metal catalysts whose lifetime has been enhanced by alloying of the active metals. 
  • Second, through the development of catalysts to produce fully formulated sustainable aviation fuel from fats oils and greases by tuning the type and strength of catalyst acid sites while maintaining the active metal sites needed for deoxygenation. 
  • Finally, the Center for Applied Energy Research has investigated the Fischer-Tropsch Synthesis for more than three decades. The synthesis is a process through which syngas (carbon monoxide and hydrogen) is converted to hydrocarbons over either iron or cobalt based catalysts, with products ranging from methane to long chain waxes. When gasified biomass or CO2 is used as the carbon source, renewable hydrocarbon fuels can be produced. 

Equipped with an experimental database going back decades, the adoption of machine learning to guide catalyst development will greatly enhance the understanding of how these catalysts function while also dramatically accelerating the speed of catalyst development.

Photo of Dr. Robert B. PaceBiography: Dr. Robert Pace first joined UK as a temporary technician in the Biofuels and Environmental Catalysis group at the Center for Applied Energy Research in 2011. He was then hired into a permanent position as assistant research scientist and continued to work at CAER on various projects, including the decarboxylation/decarbonylation of waste lipids to fuel-like hydrocarbons, the fractionation of lignocellulosic and algal biomass to produce value-added products and the use of microalgae to capture power-plant CO2 emissions. 

He subsequently joined UK’s chemistry graduate program in 2016, performing his doctoral research on the passive adsorption of NOx emitted during cold starts of automotive engines at the UK CAER under the guidance of Dr. Mark Crocker and Dr. Jack Selegue. After obtaining his Ph.D. in 2020, he returned to CAER as an Associate Research scientist, and within two years was promoted to Senior Research Scientist. Since completing his graduate degree, he has worked with the Biofuels Group; Clean Fuels and Chemicals Group; and Carbon Materials group. He also has helped to oversee the unification of the two CAER catalysis groups into the newly established Sustainable and Alternative Fuels group.

His current research focuses on the application of heterogeneous catalysis to the production of renewable fuels and chemicals with emphasis on the upgrading of waste oils to drop-in hydrocarbon fuels and the use of CO2 to produce synthetic fuels using the Fischer-Tropsch process.

Date:
Location:
CP-114

Probing nuclear collective dynamics with polarized photons at HIγS

Abstract: Electromagnetic probes provide a clean and selective window into the microscopic structure of atomic nuclei. Real photons couple directly to nuclear charge and current distributions with minimal distortion from the interaction mechanism, making resonant photon scattering particularly well suited for high-resolution studies of electric and magnetic dipole excitations. In this seminar, I will present recent photon-scattering measurements performed with quasi-monochromatic, highly polarized beams at the High Intensity Gamma-ray Source (HIγS) at the Triangle Universities Nuclear Laboratory. 

These experiments map dipole-strength distributions from low excitation energies to the vicinity of the particle-emission threshold, where the interplay of single-particle configurations and collective dynamics produces fragmented and often complex excitation patterns. Polarization asymmetries and angular distributions are used to determine multipole character, while measured intensities and decay branches provide access to reduced transition probabilities, branching ratios, and photon-strength functions. 

Selected results will be compared with state-of-the-art microscopic calculations and complementary measurements using hadronic probes. These comparisons elucidate the structure and evolution of low-energy dipole collectivity, test modern nuclear models, and constrain electromagnetic response functions relevant to nuclear reaction theory, astrophysical reaction rates, and nucleosynthesis.

Dr. Akaa Ayangeakaa

Bio: Akaa D. Ayangeakaa is an associate professor in the Department of Physics and Astronomy at the University of North Carolina at Chapel Hill and a faculty member at the Triangle Universities Nuclear Laboratory. He joined UNC in 2020 after serving as an assistant professor in the Department of Physics at the U.S.  Naval Academy in Annapolis, Maryland. Before his faculty appointments, he was a postdoctoral research fellow in the Physics Division at Argonne National Laboratory. He earned his Ph.D. in experimental nuclear physics from the University of Notre Dame, where he also completed an M.S. in experimental nuclear physics. He also holds an M.Sc. in radiation and environmental protection from the University of Surrey in the United Kingdom and a B.Sc. in physics from Benue State University.

Ayangeakaa is an experimental nuclear physicist whose research focuses on the structure and dynamics of atomic nuclei. His work investigates how the interactions of protons and neutrons give rise to collective nuclear phenomena, including shell evolution, shape coexistence, triaxial deformation, quadrupole collectivity, high-spin structure and electromagnetic response. His research group uses advanced experimental techniques such as gamma-ray spectroscopy, Coulomb excitation, nuclear resonance fluorescence and in-beam spectroscopy at major national user facilities, including TUNL, Argonne National Laboratory and the Facility for Rare Isotope Beams.

At UNC, Ayangeakaa teaches a range of undergraduate and graduate courses. His teaching portfolio includes introductory physics, experimental physics laboratory courses and classical mechanics. He is teaching PHYS 885, the introductory graduate seminar, which helps incoming graduate students develop the professional, research and academic skills needed for success in graduate school and beyond.

In addition to his teaching and research, Ayangeakaa serves as associate chair for graduate studies and director of graduate studies, where he helps oversee graduate education, student progress, admissions, mentoring and graduate-program development. He has also contributed to departmental committees related to graduate admissions, diversity, undergraduate affairs and student training.

Ayangeakaa is active in the broader nuclear physics community through national and international service. He has served on organizing and advisory committees, reviewed proposals for federal funding agencies, refereed manuscripts for leading nuclear physics journals and contributed to community-wide efforts aimed at strengthening nuclear science. Through his research, teaching, mentoring and service, he is committed to advancing the understanding of the atomic nucleus while training the next generation of scientists.

Date:
-
Location:
CP-114

Uncovering and Reprogramming Nature’s Enzymatic Chemistry

Abstract: Natural products arise from enzymes that perform remarkable chemical transformations with exquisite selectivity. Our laboratory seeks both to uncover these transformations in biological pathways and to reprogram their underlying catalytic logic for synthetic applications. 

In this seminar, I will describe our efforts in elucidating the biosynthetic pathway of structurally complex natural products, including terpenoids and izidine-type alkaloids. These studies illustrate how pathway organization, enzyme stereoselectivity and protective modifications work together to control molecular structure and biological function. 

I will also discuss our efforts to repurpose enzymes as programmable biocatalysts. By combining mechanistic insight with protein engineering, we have expanded the native reactivity of these enzymes to enable stereoselective bond formation and the construction of valuable cyclic scaffolds. Together, this work demonstrates how deciphering nature’s catalytic solutions can uncover new biochemical principles while providing a foundation for developing selective and sustainable synthetic methods.

Bio: Dr. Yang Hai received his B.S. in chemistry from Peking University in 2011 and his Ph.D. in chemistry from the University of Pennsylvania in 2016 under the guidance of David W. Christianson. His doctoral research focused on the structure-function relationships of metallohydrolases, primarily using protein X-ray crystallography. From 2016 to 2020, he conducted postdoctoral research with Yi Tang at UCLA, where he studied fungal natural product biosynthesis and discovered new enzymes and metabolic pathways through genome mining. In 2020, Hai began his independent career at the University of California, Santa Barbara. His research group investigates the biosynthesis of natural products and reprograms enzymes for applications in biocatalysis. His honors include an NIH MIRA grant, an NSF CAREER Award and an ACS Petroleum Research Fund Doctoral New Investigator Award.

Photo of Dr. Yang Hai

Date:
-
Location:
CP-114

Using Unnatural Amino Acids to Probe the Main-chain Chemistry of Neurotransmitter-activation in GABAA Receptors.

Abstract: GABAA receptors are pentameric ligand-gated ion channels (pLGICs) essential for inhibitory neurotransmission in the brain. Although their three-dimensional structures are increasingly well defined, the molecular mechanisms that couple neurotransmitter or drug binding to opening of the ion-conducting pore remain poorly understood. Here, we use both conventional and unnatural mutagenesis to demonstrate that main-chain hydrogen bonds (H-bonds) within transduction loops are crucial for transducing GABA binding to ion conduction. 

We identify an essential H-bond that couples the extracellular neurotransmitter-binding and transmembrane domains that has been conserved in inhibitory pLGICs for billions of years. Disruption of this H-bond provides a mechanistic explanation for neurological disorders including epilepsy, neurodevelopmental disability, and hyperekplexia (startle disease). These findings reveal a previously underappreciated role for main-chain chemistry in pLGIC function.

Dr. Marcel Goldschen-OhmBiography: Marcel Goldschen-Ohm’s training integrates physics, biophysics and ion channel physiology. After working in high-energy physics and quantum computing, he followed a long-standing interest in neuroscience to study ion channels. He completed his Ph.D. with Mathew Jones, where he developed expertise in GABAA receptor biophysics, patch-clamp electrophysiology and Markov modeling. As a postdoc with Baron Chanda, he developed a fluorescence-based approach for visualizing ligand binding dynamics at single molecules. At UT Austin, Marcel’s research program defines fundamental mechanisms of ion channel gating and drug modulation and its dysfunction in neurological disease.


Representation of GABA ReceptorsRepresentation of H-bond

Date:
Location:
CP-114

Doctoral Exit Seminar: Student Familiarity with the Periodic Table of the Elements: Results from Cued-Recall and Eye-Tracking Assessments on Memory

Photo of Victor OkuoAbstract: Learning element symbol-name relationships and the spatial organization of elements on the periodic table is a foundational step in learning chemistry, supporting later understanding of chemical formulas, equations, bonding and stoichiometry. Although students often rely on memorization strategies to learn periodic table content, this task is challenging due to the large number of elements and the apparent ambiguity in matching some element symbols to their names. 

This study explores students’ recall of element names when given element symbols as cues and their knowledge of element locations on the periodic table when given element names as cues. The study also examines how these two pieces vary across course groups. Seven instructional groups representing increasing levels of chemistry coursework were examined using two complementary but distinct experimental approaches: a survey-based cued-recall task assessing percentage-correct symbol-name recall, and a separate eye-tracking experiment assessing how students visually searched for selected elements on the periodic table.

The overall results with all elements considered together reveal that recall of element symbol-name relationships and visual search efficiency improve with increasing chemistry exposure, but do not follow a strictly stepwise progression. Element-specific analyses reveal that recall of symbol-name relationships developed unevenly across the periodic table: 

  • Some elements consolidate early.
  • Others strengthen gradually with increased exposure to chemistry.
  • A subset remain weakly recalled even among upper-level undergraduate and graduate students. 

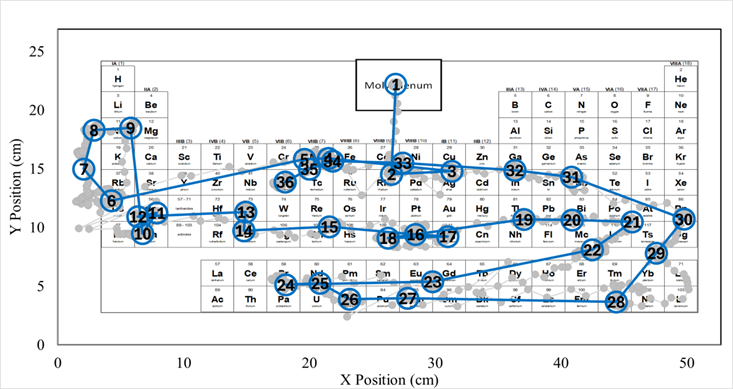
Visual search efficiency follows a similar pattern, shifting from exploratory search to shorter, more focused search paths in advanced course groups. Recall and visual search efficiency were similar but not equivalent. In some cases, elements were located efficiently despite weak or absent recall, whereas in others  elements recalled were associated with inefficient search behavior. 

These findings indicate that symbol-name knowledge and spatial knowledge of element locations constitute distinct but interacting memory components. At the upper levels of instruction, there seems to be an integration of these components, enabling students to use the periodic table more effectively as a structured representation rather than relying solely on element symbol-name associations.

Illustration of the periodic table

KEYWORDS: Periodic table, symbol–name relationalships, visual search, search behavior, consolidation, integration

Date:
Location:
CP-114

Doctoral Exit Seminar: Student Familiarity with the Periodic Table of the Elements: Results from Cued-Recall and Eye-Tracking Assessments on Memory

Photo of Victor OkuoAbstract: Learning element symbol-name relationships and the spatial organization of elements on the periodic table is a foundational step in learning chemistry, supporting later understanding of chemical formulas, equations, bonding and stoichiometry. Although students often rely on memorization strategies to learn periodic table content, this task is challenging due to the large number of elements and the apparent ambiguity in matching some element symbols to their names. 

This study explores students’ recall of element names when given element symbols as cues and their knowledge of element locations on the periodic table when given element names as cues. The study also examines how these two pieces vary across course groups. Seven instructional groups representing increasing levels of chemistry coursework were examined using two complementary but distinct experimental approaches: a survey-based cued-recall task assessing percentage-correct symbol-name recall, and a separate eye-tracking experiment assessing how students visually searched for selected elements on the periodic table.

The overall results with all elements considered together reveal that recall of element symbol-name relationships and visual search efficiency improve with increasing chemistry exposure, but do not follow a strictly stepwise progression. Element-specific analyses reveal that recall of symbol-name relationships developed unevenly across the periodic table: 

  • Some elements consolidate early.
  • Others strengthen gradually with increased exposure to chemistry.
  • A subset remain weakly recalled even among upper-level undergraduate and graduate students. 

Visual search efficiency follows a similar pattern, shifting from exploratory search to shorter, more focused search paths in advanced course groups. Recall and visual search efficiency were similar but not equivalent. In some cases, elements were located efficiently despite weak or absent recall, whereas in others  elements recalled were associated with inefficient search behavior. 

These findings indicate that symbol-name knowledge and spatial knowledge of element locations constitute distinct but interacting memory components. At the upper levels of instruction, there seems to be an integration of these components, enabling students to use the periodic table more effectively as a structured representation rather than relying solely on element symbol-name associations.

Illustration of the periodic table

KEYWORDS: Periodic table, symbol–name relationalships, visual search, search behavior, consolidation, integration

Date:
Location:
CP-114

Doctoral Exit Seminar: Targeting Intrinsically Disordered Proteins with Ligand-Guided Gold(III)-Mediated Covalent Chemistry

Photo of Udara Munugoda HewageAbstract: The human genome encodes approximately 20,000 proteins, of which about 85% lack well-defined druggable binding pockets, leaving most of the disease-relevant proteins “undruggable."

Small molecule chemical probes are essential for modulating protein function and serve as leads for therapeutic development. However, the majority of the human proteome remains inaccessible to conventional drug-like molecules due to the absence of suitable binding pockets. Intrinsically disordered proteins (IDPs) and intrinsically disordered regions (IDRs) represent a particularly hard-to-drug class within this undruggable proteome. Although IDPs lack stable tertiary structures, they still play central roles in transcriptional regulation, oncogenic signaling, and cellular stress responses. 

Mass spectrometry-based cysteine chemoproteomics has emerged as a promising approach to address this druggability gap by mapping cysteine reactivity across the proteome. Cysteine-thiol (Cys-SH) is the preferred nucleophile for site-selective protein modification because of its high intrinsic reactivity, low natural abundance, and regulatory significance. Nonetheless, significant challenges remain, as conventional cysteine-targeted electrophiles (warheads) predominantly modify solvent-exposed residues in well-folded protein domains, leaving IDPs and IDRs largely inaccessible to covalent targeting. Existing strategies for engaging IDPs often depend on serendipitous or cryptic pocket targeting, which do not provide a generalizable or mechanistically understood framework for capturing the disordered cysteinome.

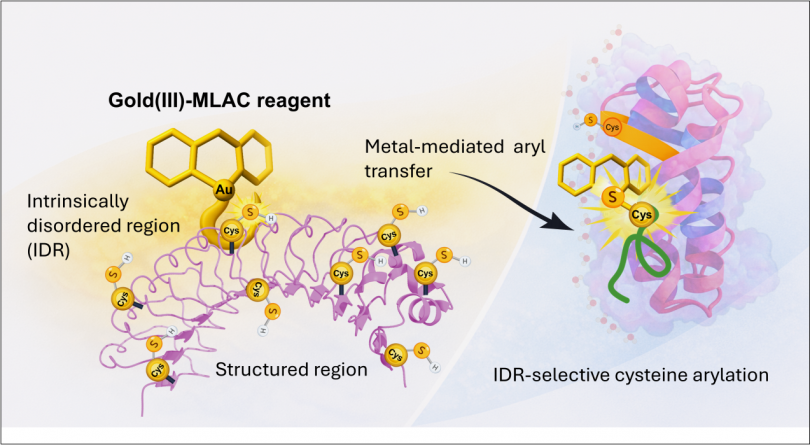
This work spans multiple disciplines by integrating structure-guided design, chemical synthesis, analytical characterization, quantitative mass spectrometry-based chemoproteomics, computational modeling, structural data integration and cell-based validation to address these challenges. We hypothesized that cyclometalated Au(III)[C^N] frameworks bearing unconventional and tunable bulky monodentate phosphine ancillary ligands would modulate steric shielding at the gold(III) center to enable site-selective cysteine arylation within disordered protein regions. 

These gold(III) complexes form irreversible C(sp²)-heteroatom bonds via metal-mediated aryl transfer to Cys-SH, and their square-planar geometry and relativistic stabilization enhance their electrophilicity and affinity for soft nucleophiles like Cys-SH. The overall outcome is a new class of gold(III) arylating reagents with tunable ancillary ligands that covalently target cysteines within dynamically unstable IDRs, thereby increasing local protein structural stability while preserving existing disulfide bonds. Altogether, we designed a next-generation biorthogonal cyclometalated gold(III) probe platform and expanded our in-house technique, Metal-mediated Ligand Affinity Chemistry (MLAC), by conjugating protein-binding ligands to gold(III) arylating reagents for proximity-driven, site-selective covalent modification of native IDPs.

Using this platform, we demonstrated IDR-selective covalent targeting of historically undruggable proteins, including redox-sensitive Galectin-1, disordered regions of Heme Oxygenase-2 (HMOX2), and mutant KRAS G12C, achieving a "speed + IDP selectivity" profile unattainable by conventional electrophiles. We profiled the human cysteinome in triple-negative breast cancer (TNBC) cell lines through gold(III)-mediated one-pot CuAAC biorthogonal click chemistry coupled with quantitative chemoproteomics. This approach identified 391 IDPs, including 261 undruggable proteins that are not targeted by FDA-approved drugs or small molecules listed in DrugBank or ChEMBL, and deposited the complete dataset in the PRIDE proteomics repository.

However, the impact of our MLAC work will be limited if researchers cannot apply this platform in biologically relevant systems. Building on these chemoproteomic findings, we then developed gold(III) arylating reagents conjugated to Lenalidomide to selectively target cysteines coordinated to the zinc(II) metal center in Cereblon, thereby reprogramming the specificity of E3 ligases and expanding the degradable proteome. We further extended this chemistry towards therapeutic translation to engineer site-specific antibody-drug conjugates (ADCs) with enhanced homogeneity and drug-to-antibody ratio (DAR) while preserving antibody activity through cysteine-selective bioconjugation.

Collectively, this dissertation expands our understanding of the druggable cysteinome by establishing cyclometalated gold(III) chemistry as a unified framework that connects IDP targeting, quantitative chemoproteomic mapping, organometallic protein degradation, and precision bioconjugation. This work delivers structure-guided principles for gold(III) probe design and mass spectrometry-based workflows that enable covalent targeting of proteins previously considered undruggable, thereby deciphering the mechanisms underlying protein undruggability to advance covalent drug development and translational cancer therapeutics.

Illustration of Gold(III)-MLAC reagent

Date:
Location:
CP-114

Doctoral Exit Seminar: Targeting Intrinsically Disordered Proteins with Ligand-Guided Gold(III)-Mediated Covalent Chemistry

Photo of Udara Munugoda HewageAbstract: The human genome encodes approximately 20,000 proteins, of which about 85% lack well-defined druggable binding pockets, leaving most of the disease-relevant proteins “undruggable."

Small molecule chemical probes are essential for modulating protein function and serve as leads for therapeutic development. However, the majority of the human proteome remains inaccessible to conventional drug-like molecules due to the absence of suitable binding pockets. Intrinsically disordered proteins (IDPs) and intrinsically disordered regions (IDRs) represent a particularly hard-to-drug class within this undruggable proteome. Although IDPs lack stable tertiary structures, they still play central roles in transcriptional regulation, oncogenic signaling, and cellular stress responses. 

Mass spectrometry-based cysteine chemoproteomics has emerged as a promising approach to address this druggability gap by mapping cysteine reactivity across the proteome. Cysteine-thiol (Cys-SH) is the preferred nucleophile for site-selective protein modification because of its high intrinsic reactivity, low natural abundance, and regulatory significance. Nonetheless, significant challenges remain, as conventional cysteine-targeted electrophiles (warheads) predominantly modify solvent-exposed residues in well-folded protein domains, leaving IDPs and IDRs largely inaccessible to covalent targeting. Existing strategies for engaging IDPs often depend on serendipitous or cryptic pocket targeting, which do not provide a generalizable or mechanistically understood framework for capturing the disordered cysteinome.

This work spans multiple disciplines by integrating structure-guided design, chemical synthesis, analytical characterization, quantitative mass spectrometry-based chemoproteomics, computational modeling, structural data integration and cell-based validation to address these challenges. We hypothesized that cyclometalated Au(III)[C^N] frameworks bearing unconventional and tunable bulky monodentate phosphine ancillary ligands would modulate steric shielding at the gold(III) center to enable site-selective cysteine arylation within disordered protein regions. 

These gold(III) complexes form irreversible C(sp²)-heteroatom bonds via metal-mediated aryl transfer to Cys-SH, and their square-planar geometry and relativistic stabilization enhance their electrophilicity and affinity for soft nucleophiles like Cys-SH. The overall outcome is a new class of gold(III) arylating reagents with tunable ancillary ligands that covalently target cysteines within dynamically unstable IDRs, thereby increasing local protein structural stability while preserving existing disulfide bonds. Altogether, we designed a next-generation biorthogonal cyclometalated gold(III) probe platform and expanded our in-house technique, Metal-mediated Ligand Affinity Chemistry (MLAC), by conjugating protein-binding ligands to gold(III) arylating reagents for proximity-driven, site-selective covalent modification of native IDPs.

Using this platform, we demonstrated IDR-selective covalent targeting of historically undruggable proteins, including redox-sensitive Galectin-1, disordered regions of Heme Oxygenase-2 (HMOX2), and mutant KRAS G12C, achieving a "speed + IDP selectivity" profile unattainable by conventional electrophiles. We profiled the human cysteinome in triple-negative breast cancer (TNBC) cell lines through gold(III)-mediated one-pot CuAAC biorthogonal click chemistry coupled with quantitative chemoproteomics. This approach identified 391 IDPs, including 261 undruggable proteins that are not targeted by FDA-approved drugs or small molecules listed in DrugBank or ChEMBL, and deposited the complete dataset in the PRIDE proteomics repository.

However, the impact of our MLAC work will be limited if researchers cannot apply this platform in biologically relevant systems. Building on these chemoproteomic findings, we then developed gold(III) arylating reagents conjugated to Lenalidomide to selectively target cysteines coordinated to the zinc(II) metal center in Cereblon, thereby reprogramming the specificity of E3 ligases and expanding the degradable proteome. We further extended this chemistry towards therapeutic translation to engineer site-specific antibody-drug conjugates (ADCs) with enhanced homogeneity and drug-to-antibody ratio (DAR) while preserving antibody activity through cysteine-selective bioconjugation.

Collectively, this dissertation expands our understanding of the druggable cysteinome by establishing cyclometalated gold(III) chemistry as a unified framework that connects IDP targeting, quantitative chemoproteomic mapping, organometallic protein degradation, and precision bioconjugation. This work delivers structure-guided principles for gold(III) probe design and mass spectrometry-based workflows that enable covalent targeting of proteins previously considered undruggable, thereby deciphering the mechanisms underlying protein undruggability to advance covalent drug development and translational cancer therapeutics.

Illustration of Gold(III)-MLAC reagent

Date:
Location:
CP-114

Doctoral Exit Seminar: Chemical Modulation of MYC and Mitochondrial Mechanisms in Biology

Photo of Oluwatosin ObisesanAbstract: Master regulatory control programs coordinate genetic activity, metabolic state and stress adaptation across cellular systems. Targeting these programs remains a major challenge in chemical biology due to structural complexity, network redundancy, and context-dependent regulation. Understanding how chemical tools can reprogram interwoven regulatory layers is essential for overcoming limitations of current therapeutic strategies. 

This dissertation examines how small molecules can modulate two interconnected master regulators: the transcription factor MYC and the mitochondrion. MYC is a protein that regulates the transcription of about 15% of genes in the body, while the mitochondrion is the central organelle governing bioenergetic and redox homeostasis, both of which are frequently altered in inflammatory states.

MYC drives numerous human cancers and represents a viable yet historically intractable target due to its intrinsically disordered structure and lack of a defined binding pocket. Many MYC inhibitors fail because reversible interactions are insufficient to stabilize engagement of this unstable protein.

 To address this challenge, I developed multiple strategies to achieve functional chemical engagement of MYC. An in-house platform termed Metal-mediated Affinity Chemistry enabled proximity-guided, site-selective targeting of MYC. The known MYC-MAX disruptor 10058-F4 was conjugated to an Au(III)-based warhead capable of chemo-selective cysteine modification within intrinsically disordered regions, stabilizing small-molecule-MYC interaction and enhancing potency without reliance on conventional pocket binding.

In parallel, through the development of MY05, I established direct intracellular inhibition of MYC. MY05 selectively disrupts MYC-MAX heterodimerization and attenuates MYC-dependent transcriptional programs in cancer models, demonstrating that intrinsically disordered transcription factors can be chemically engaged with meaningful functional consequences. MY05 also provides a chemical framework for the subsequent development of potent covalent modifiers and degraders of MYC.

To extend beyond direct inhibition, complementary approaches were pursued to regulate MYC indirectly through its upstream biochemical control. Targeting HMOX2 revealed a chemical axis that promotes MYC depletion while simultaneously modulating mitochondrial function and redox balance. These findings uncover a mechanistic interface between mitochondrial metabolism and oncogenic transcription, demonstrating that bioenergetic state influences MYC-dependent gene expression. 

Chemical perturbation of mitochondrial function was shown to reprogram metabolic states and selectively challenge disease-associated phenotypes. Mitochondria thus function as regulatory nexuses capable of reshaping proliferative and inflammatory signaling networks.

Building on this intersection, direct modulation of mitochondrial regulatory programs was investigated in inflammatory contexts. A brain-penetrant Au(III)-based compound, AuPhos, induced mitochondrial biogenesis, enhanced oxidative capacity, regulated inflammatory signaling and promoted coordinated transcriptional remodeling. In a model of traumatic brain injury, mitochondrial enhancement supported molecular programs associated with metabolic resilience and tissue repair.

Collectively, this work establishes a framework for chemical reprogramming of transcriptional and bioenergetic control systems. Through small-molecule MYC engagement, redox-mediated MYC regulation, and mitochondria-driven transcriptional remodeling, these studies define the functional interplay among master regulatory layers. Further elucidation of the MYC-mitochondrial interface may enable precision modulation of cellular state and inform the development of next-generation therapeutics.

Depictions of development of mitochondria-targeting small-molecules, and regulating the c-MYC oncoprotein.

Date:
Location:
CP-114